 ultimate-guide
Understanding cGMP Standards for Research Peptides
Table of Contents
- What cGMP Standards Mean for Research Peptides
- cGMP vs GMP: Understanding the Difference
- FDA Regulations and the Role of Current Good Manufacturing Practice
- Peptide Quality Control Testing: Methods and Standards
- Reading and Interpreting Your Peptide Certificate of Analysis
- cGMP Peptide Manufacturing: Process Controls and Documentation
- What cGMP Does Not Certify: Clarifying Research-Use Status
- How to Verify a Supplier's cGMP Claims
- Frequently Asked Questions
Last Updated: October 13, 2026
What cGMP Standards Mean for Research Peptides
Current Good Manufacturing Practice (cGMP) standards represent a set of regulatory requirements that govern how peptides are manufactured, tested, and documented to ensure quality and consistency. At Everform Research, we understand that cGMP compliance is foundational to delivering peptides you can trust for reproducible research outcomes.
cGMP standards establish minimum requirements for facilities, equipment, personnel, processes, and quality systems. They're not a certification that a peptide is safe for human use, rather, they're a framework ensuring that whatever is claimed on the label is actually in the vial, batch after batch. The standards cover everything from raw material sourcing through final product release, including documentation that tracks every step of manufacture.
For research peptides, understanding cGMP standards for research peptides means recognising the difference between a supplier who follows good practices and one whose processes are verifiable, auditable, and aligned with regulatory expectations. This distinction matters when you're designing experiments where consistency across batches determines whether your data is reproducible or compromised.
cGMP vs GMP: Understanding the Difference
The distinction between GMP and cGMP is subtle but important. Good Manufacturing Practice (GMP) refers to the broader set of guidelines and best practices for pharmaceutical and chemical manufacturing. Current Good Manufacturing Practice (cGMP) is the specific, enforceable regulatory standard maintained by the Food and Drug Administration (FDA) that applies to drug manufacturing. The word "current" is doing real work here: it signals that the expectations are updated as science, technology, and manufacturing knowledge advance, so a facility following cGMP is following the FDA's present-day expectations rather than a frozen, outdated version.
The clearest way to see the difference is to look at what each term actually obliges a manufacturer to do.
- GMP describes a family of good-practice frameworks. It can refer to international guidance, industry consensus standards, or a manufacturer's own internal quality programme. It is a useful signal, but it is not a single enforceable rulebook with a named regulator behind it.
- cGMP is the enforceable standard. In the United States, drug manufacturing requirements sit in Title 21 of the Code of Federal Regulations, Part 211 (21 CFR 211), which covers personnel, facilities, equipment, process controls, quality assurance, and record-keeping. A facility claiming cGMP compliance is claiming alignment with that framework.
The practical difference for a researcher is accountability. A cGMP facility operates under a regulatory framework with inspection protocols, deviation management systems, and documented corrective actions. When something goes wrong, there is a paper trail. A facility that describes itself only as "GMP" may have excellent practices, but without the regulatory overlay there is less external verification that those practices are consistently applied.
It also helps to understand how the FDA actually enforces cGMP, because that is what gives the standard its teeth. The FDA inspects manufacturing facilities, issues warning letters, and can take enforcement action when requirements are not met. This creates an incentive structure: a cGMP-compliant facility has invested in systems and documentation specifically to withstand that scrutiny. When you source from a cGMP manufacturer, you are sourcing from an operation that has demonstrated it can meet federal standards or face consequences.
For research peptides specifically, the picture is more nuanced than a simple GMP-versus-cGMP comparison suggests. FDA oversight applies differently depending on the intended use of the peptide. Peptides manufactured under cGMP for clinical drug development face full FDA scrutiny. Research-grade peptides, by contrast, operate in a different regulatory space: they are not subject to FDA pre-market approval, but a supplier claiming cGMP compliance is still expected to follow the manufacturing standards themselves. That is the gap where careful reading pays off, a cGMP claim tells you about the manufacturing system, not about the regulatory status of the product in your hands.
A useful mental model: GMP tells you a manufacturer intends to follow good practice; cGMP tells you which rulebook, which regulator, and which enforcement mechanism stand behind that intent. For reproducible research, the second is the one you can actually verify.
FDA Regulations and the Role of Current Good Manufacturing Practice
The FDA establishes cGMP regulations under Title 21 of the Code of Federal Regulations, Part 211 (21 CFR 211), which outlines requirements for pharmaceutical manufacturing. These regulations cover personnel qualifications, facility design, equipment validation, process controls, quality assurance, and record-keeping. FDA guidance on pharmaceutical cGMP regulations provides the authoritative framework that manufacturers must follow.
Understanding cGMP standards for research peptides requires recognising that FDA oversight applies differently depending on the intended use of the peptide. Peptides manufactured under cGMP for clinical drug development face full FDA scrutiny. Research-grade peptides, by contrast, operate in a different regulatory space, they're not subject to FDA pre-market approval, but suppliers claiming cGMP compliance are still expected to follow the manufacturing standards themselves.
The FDA's role is enforcement and inspection. Facilities manufacturing under cGMP standards are subject to FDA inspections, warning letters, and enforcement actions if they fail to meet requirements. This regulatory pressure creates an incentive structure: a cGMP-compliant facility has invested in systems and documentation specifically to pass FDA scrutiny. When you source from a cGMP manufacturer, you're sourcing from an operation that has demonstrated it can meet federal standards or face consequences.
Peptide Quality Control Testing: Methods and Standards
Quality control (QC) testing is where cGMP standards translate into tangible assurance. A comprehensive QC programme for peptides includes testing for identity, purity, potency, sterility, endotoxin levels, and stability. According to GenScript's guidance on GMP compliance for peptide quality control, these tests support both product release decisions and shelf-life claims.
The purpose of QC testing is twofold: verify that the batch meets specifications and identify any deviations early enough to prevent release of out-of-specification material. Each test method has defined acceptance criteria, and results are documented in a Certificate of Analysis (COA) that accompanies the product.
Identity, Purity, and Potency Assessment
Identity testing confirms that the peptide is what you ordered, the correct amino acid sequence and molecular weight. Mass spectrometry is the standard method, providing a precise molecular weight that matches the theoretical value for your peptide. If the mass is off, either the wrong peptide was synthesised or contamination occurred during manufacturing.
Purity testing measures the percentage of the desired peptide versus impurities. High-performance liquid chromatography (HPLC) is the most common method, separating the peptide from related compounds and quantifying each component. A purity specification might be ≥95%, meaning at least 95% of the material is the target peptide. The remaining <5% could be truncated sequences, oxidised forms, or other related peptides generated during synthesis.
Potency testing assesses biological activity when relevant. For some research applications, the amino acid sequence alone isn't sufficient, the peptide must also exhibit expected biological function. Potency assays are application-specific and may involve cell-based assays or biochemical activity measurements.
Sterility, Endotoxin, and Contamination Testing
Sterility testing detects bacterial and fungal contamination. For lyophilised (freeze-dried) peptides, sterility testing is typically performed on the final product. The test involves incubating the peptide in growth media and observing for microbial growth over a defined period.
Endotoxin testing measures lipopolysaccharides from gram-negative bacteria. Even if bacteria are killed during manufacturing, their endotoxin can remain and trigger immune responses. Endotoxin limits are specified in parts per million (ppm) or endotoxin units per vial, depending on the intended use.
Contamination control extends beyond testing. It begins with raw material sourcing, continues through manufacturing in controlled environments (cleanrooms), and includes environmental monitoring during production. According to BOC Sciences guidance on cGMP peptide manufacturing controls, environmental monitoring programmes track viable and non-viable particles to detect trends that might indicate process drift.
Reading and Interpreting Your Peptide Certificate of Analysis
A Certificate of Analysis (COA) is the quality record for a specific batch. It documents the results of all QC testing and confirms that the batch meets specifications before release. Learning to read a COA is essential for verifying supplier claims and troubleshooting if results don't match expectations.
A typical COA includes batch number, manufacturing date, expiry date, and a table of test results. Each test has a specification (the acceptable range) and the actual result. If the result falls within the specification, it's marked as "Pass" or "Compliant." If it's outside the range, it's flagged as "Out of Specification" or "OOS," which means the batch should not have been released.
Look for the testing methods used. HPLC-UV for purity, LC-MS for identity, and the specific assay method for potency should all be named. The COA should also identify the testing laboratory. If testing was performed in-house by the manufacturer, that's less independent verification than third-party testing. Everform Research emphasises third-party testing to provide you with transparent, unbiased quality data that you can trust.
The COA should include a signature or electronic approval from the QA department, indicating that someone with authority has reviewed the results and authorised release. This audit trail is part of cGMP compliance, it creates accountability for the decision to release the batch.
cGMP Peptide Manufacturing: Process Controls and Documentation
cGMP peptide manufacturing is built on process controls and comprehensive documentation. Every step from raw material receipt through final packaging is documented. Process controls ensure that manufacturing happens consistently, producing batches with predictable quality.
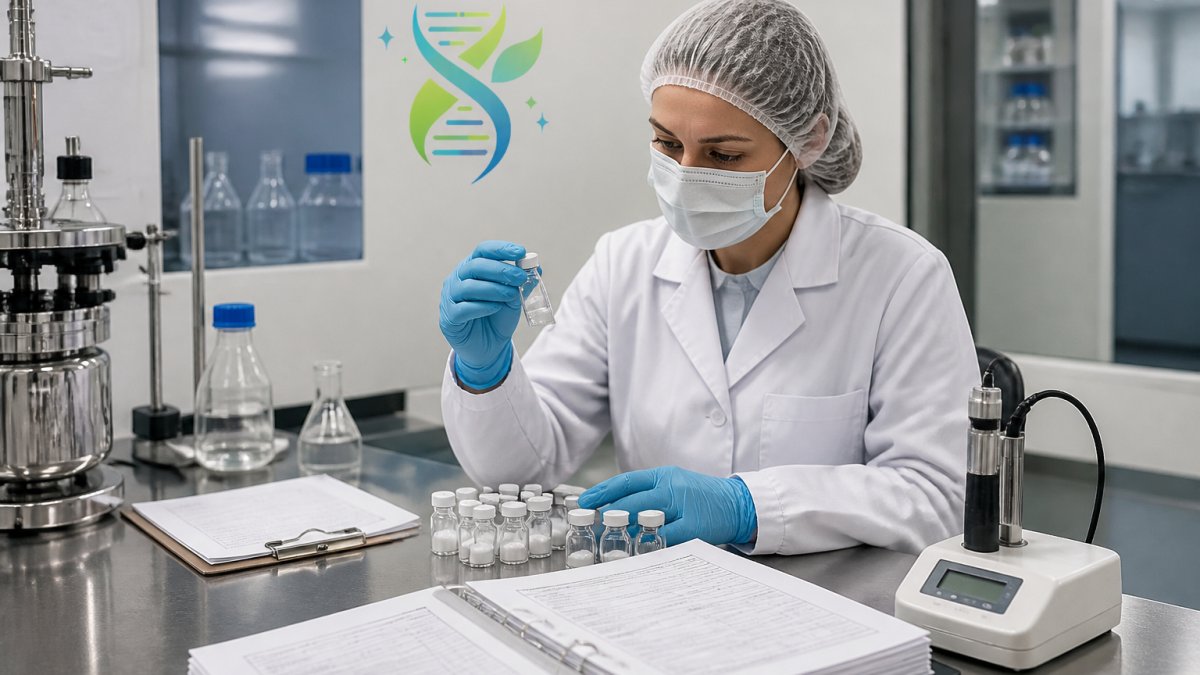
Process controls include in-process tests that monitor quality during manufacturing, not just at the end. For peptide synthesis, in-process controls might include monitoring pH, temperature, and reaction completion at defined stages. If a parameter drifts outside acceptable limits, the batch is stopped and investigated rather than allowed to continue to completion.
Raw Material Traceability and Supplier Qualification
cGMP requires that all raw materials, amino acids, coupling reagents, solvents, are qualified before use. Supplier qualification means the manufacturer has evaluated the supplier's capability to provide consistent, documented material. Raw materials must come with Certificates of Analysis from the supplier, and the receiving organisation must verify that materials meet specifications before they're used in production.
Material traceability is a cornerstone of cGMP. Every batch of peptide must be traceable back to the specific lots of raw materials used. If a problem is discovered in a finished batch, traceability allows the manufacturer to identify which other batches might have been affected and initiate a recall if necessary. This traceability extends to water systems, gases, and other utilities used in manufacturing.
Batch Records and Standard Operating Procedures
A batch record is the manufacturing history of a specific batch. It documents every step taken, every test performed, every deviation encountered, and every decision made during manufacture. Batch records are the primary evidence that cGMP standards were followed. They're retained for the life of the product plus a defined period (often 5-10 years for research-grade materials).
Standard Operating Procedures (SOPs) are the written instructions that govern how manufacturing is performed. An SOP for peptide synthesis describes the starting materials, equipment to be used, step-by-step instructions, in-process tests, acceptance criteria, and documentation requirements. Every person involved in manufacturing must be trained on the relevant SOPs and must follow them consistently.
When deviations occur, a temperature excursion, an unexpected result, a missed step, they're documented in the batch record. The deviation triggers an investigation to determine root cause and whether the batch is still acceptable for release. This deviation management system is central to cGMP; it transforms problems into learning opportunities and ensures transparency about manufacturing challenges.
What cGMP Does Not Certify: Clarifying Research-Use Status
This is where clarity is essential. cGMP compliance means the manufacturing process meets regulatory standards. It does not mean the peptide is approved for human use, safe for human consumption, or suitable for clinical application. cGMP is a manufacturing standard, not a safety or efficacy claim.
A research-grade peptide manufactured under cGMP standards is still a research chemical. It may be of pharmaceutical quality, but it hasn't undergone the clinical trials, safety studies, and regulatory review required for approval as a drug. The distinction matters because it defines the appropriate use case and the level of regulatory oversight that applies.
cGMP also doesn't certify that the peptide will work for your specific application. Quality control testing confirms that the peptide meets its specification, the correct sequence, defined purity, and expected potency within the scope of available testing methods. But whether that peptide will produce the results you expect in your experiments depends on your experimental design, your protocols, and the relevance of the peptide to your research question.
How to Verify a Supplier's cGMP Claims
This section walks through a practical verification sequence, from the facility level down to the individual batch, and flags where claims commonly outrun the evidence.
Start with facility scope, not the product page. A cGMP claim should describe a specific facility and a specific scope of operations. Ask which site manufactures the peptide, what operations that site is qualified for, and whether the claim covers synthesis, purification, lyophilisation, and packaging or only some of those steps. A supplier that cannot name the site or describe its scope is making a marketing claim, not a quality claim.
Ask for audit evidence, not just certificates. Certifications such as ISO 9001 (general quality management) or ISO 13485 (quality management for medical devices) indicate that an independent auditor has reviewed the facility's systems. They do not replace cGMP compliance, but they are external verification that a quality system exists and is audited. Pair that with any inspection reports, third-party audit summaries, or customer audit outcomes the supplier is willing to share. A genuine facility usually has something to show; a reseller usually does not.
Read the Certificate of Analysis (COA) as a document, not a badge. A COA from a cGMP facility should reference specific test methods, list results against defined specifications, and carry an approval signature from the quality unit. Watch for vague language such as "meets standards" without naming the standard, or purity figures with no method attached. If the COA does not say how a number was produced, the number is not verifiable.
Understand what each analytical method can and cannot establish. This is where most verification efforts quietly fail.
- Mass spectrometry confirms molecular identity, that the mass matches the expected sequence. It does not, on its own, prove purity or biological activity.
- Chromatography (typically HPLC) quantifies purity, the percentage of the target peptide versus related impurities. It does not confirm identity, and results depend on the column, mobile phase, and detection wavelength used.
- Endotoxin testing measures lipopolysaccharides from gram-negative bacteria. It says nothing about peptide identity or potency.
- Potency assays assess biological activity, but they are application-specific and cannot be generalised across research uses.
No single method certifies a peptide. Each one answers a narrow question, and a credible COA shows you which question each test was designed to answer.
Request batch-specific records. For critical work, ask for the batch record summary or a detailed COA for the exact lot you are purchasing, not a representative example. Batch records document every step, test, deviation, and release decision for that lot. A supplier that can provide batch-specific detail is operating a real quality system; one that can only offer a generic COA is not.
Trace the raw materials. cGMP requires that amino acids, coupling reagents, and solvents be qualified before use and traceable back to specific supplier lots. Ask how the manufacturer qualifies suppliers and whether material COAs are retained. If a supplier cannot describe its supplier qualification process, the traceability chain is broken at the first link.
Check who performs the testing. In-house testing is not automatically suspect, but third-party testing removes the financial incentive to pass a marginal batch. Ask which laboratory performed each test and whether that laboratory is independent of the manufacturer.
| Verification Step | What to Look For | Red Flag |
|---|---|---|
| Facility scope | Named site, defined operations, qualified steps | No site named or scope left vague |
| Audit evidence | Inspection reports, third-party audit summaries, ISO certifications | No external verification of any kind |
| Certificate of Analysis | Named methods, defined specs, approval signature | "Meets standards" with no method or spec |
| Method understanding | Each test tied to a specific question it answers | Purity or identity claimed without a method |
| Batch records | Batch-specific COA or record summary | Only a generic, representative COA |
| Raw material traceability | Supplier qualification process, material COAs | Cannot describe sourcing or qualification |
| Testing laboratory | Independent third-party lab named | No clarity on who tested or how |
A final point worth stating plainly: cGMP compliance is a manufacturing standard, not a certification of safety, clinical suitability, or approval for human use. A research-grade peptide made under cGMP is still a research chemical. Verifying a supplier's claims tells you how carefully the material was made and documented, it does not change what the material is appropriate for.
For premium-quality peptide products backed by rigorous quality standards, transparency, and exceptional customer service, explore Everform Research's compounds. Use discount code EVER15 for 15% off your order.
Frequently Asked Questions
What does cGMP mean in peptide manufacturing?
cGMP stands for current Good Manufacturing Practice. It is a set of FDA regulations that establish minimum standards for the design, monitoring, and control of manufacturing processes and facilities. For research peptides, cGMP compliance means the manufacturer follows documented procedures for raw material testing, process controls, quality assurance, and batch record management. However, cGMP certification does not imply the peptide is approved for human use, it confirms manufacturing quality and consistency for research applications.
How is cGMP different from GMP?
GMP (Good Manufacturing Practice) refers to general quality standards for manufacturing. cGMP (current Good Manufacturing Practice) is the FDA's specific, enforceable regulatory standard that requires manufacturers to follow current scientific understanding and best practices. The word 'current' means the standards evolve as technology and knowledge advance. For peptide suppliers, cGMP compliance is more rigorous and formally documented than basic GMP, with regular audits and corrective action protocols.
Does cGMP certification guarantee peptide quality?
cGMP compliance ensures a manufacturer follows strict controls for identity, purity, potency, and batch consistency. It does not guarantee the peptide will perform in your specific experiment or application. A cGMP-manufactured peptide meets documented specifications and has undergone rigorous quality control testing, but results depend on your experimental design, storage conditions, and how the peptide is handled. Always review the certificate of analysis to confirm the batch meets your requirements.
What testing should accompany a cGMP research peptide?
A cGMP peptide should include a certificate of analysis (COA) documenting identity testing (typically mass spectrometry), purity analysis (chromatography), potency or strength measurement, and stability data. Depending on the peptide, sterility and endotoxin testing may also be included. The COA shows batch-specific results and specifications, allowing you to verify the compound meets quality standards before use. Third-party testing adds an independent verification layer beyond the manufacturer's internal quality assurance.
Why does batch-to-batch consistency matter in research peptides?
Research relies on reproducibility. If one batch is 98% pure and the next is 96%, your experimental outcomes may vary unpredictably, making it difficult to isolate the effects of your intervention from manufacturing variation. cGMP processes include in-process controls and batch release criteria to maintain tight specifications across production runs. This consistency is essential for high-throughput screening, multi-site studies, and publishing results that other laboratories can replicate.
What is a batch record and why does it matter?
A batch record is a documented history of every step in manufacturing a specific peptide lot, including raw materials used, process parameters, testing results, and approvals. It creates an audit trail that proves the batch was made according to standard operating procedures. For research institutions with compliance requirements, batch records provide the documentation needed to verify cGMP claims and demonstrate due diligence in supplier selection. They also enable traceability if a quality issue arises.
Can I use a cGMP research peptide in human studies?
cGMP compliance addresses manufacturing quality, not regulatory approval for human use. A cGMP-manufactured research peptide is not automatically approved for human clinical trials or therapeutic application. If your research progresses to human studies, additional regulatory pathways (such as FDA IND approval) are required. Always consult your institutional review board and regulatory affairs team before any human application, even if the peptide is cGMP-compliant.